Understanding the 510(k) Acceptance Review
The FDA’s Refuse to Accept (RTA) policy establishes the minimum completeness criteria a medical device manufacturer must satisfy before a 510(k) premarket notification proceeds to substantive review. It helps ensure that the submission contains the information FDA needs to evaluate substantial equivalence to a legally marketed predicate device.
The acceptance review is intended to check the presence and organization of required information, not to determine whether the submitted scientific evidence is adequate to demonstrate substantial equivalence. That assessment takes place during substantive review.
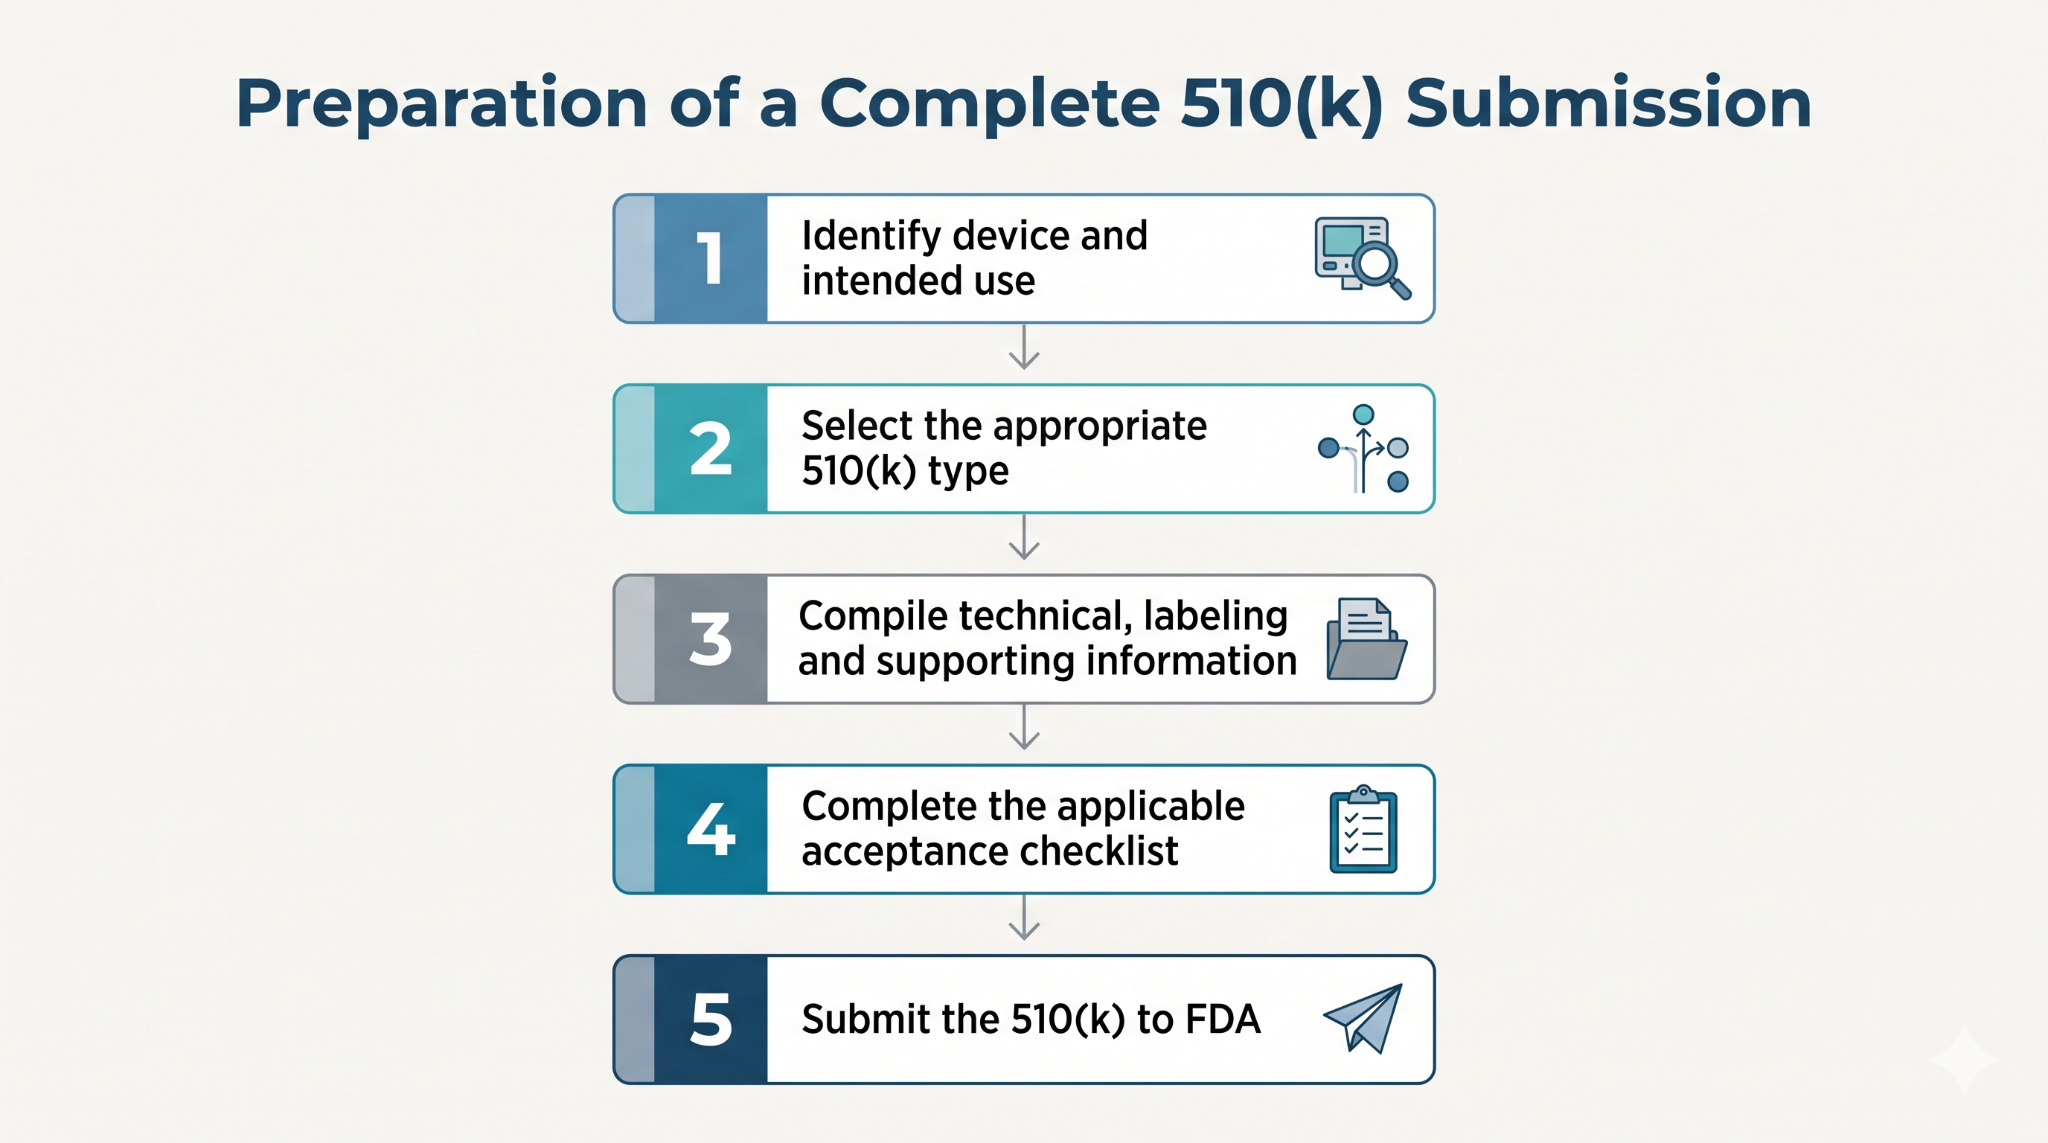
Preparing a Complete 510(k) Submission
Before submitting a 510(k), manufacturers should identify the appropriate submission type Traditional, Abbreviated, or Special and prepare the information required by the applicable acceptance checklist. The submission should describe the device, its intended use, technological characteristics, labeling, and supporting performance information.
Manufacturers should also review applicable FDA regulations, device specific guidance documents, recognized consensus standards, and special controls. The objective is to ensure that the required information is included or that a clear rationale is provided when an item is not applicable or an alternative approach is used.
A well organized submission helps FDA reviewers locate the necessary information and reduces the risk of administrative delays.
FDA Acceptance Review
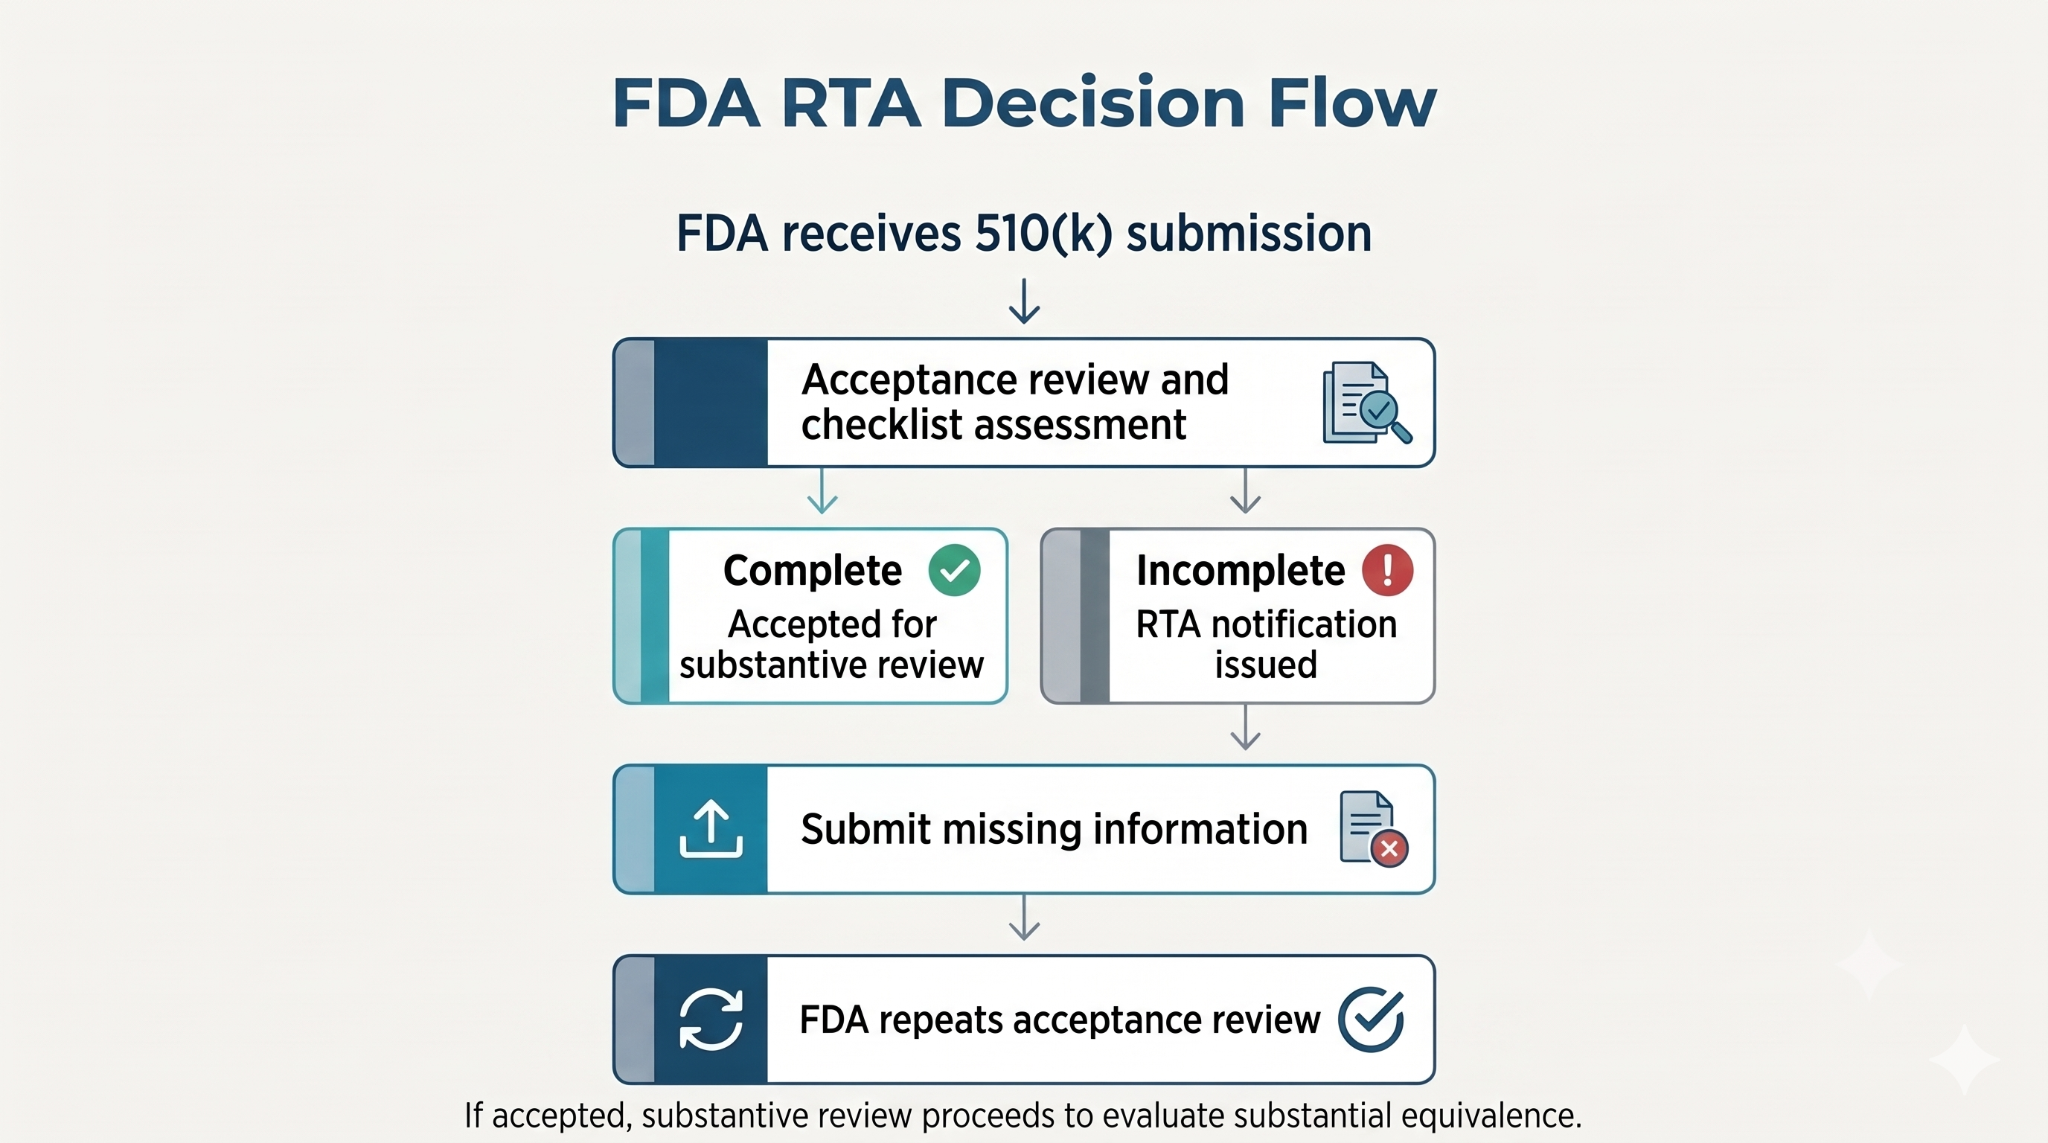
After receiving the submission, FDA conducts an acceptance review to determine whether the 510(k) is administratively complete. Under the guidance, FDA aims to complete this review within 15 calendar days of receipt.
The reviewer uses the checklist applicable to the submission type and evaluates the preliminary questions and required elements. These checks include whether the product is a medical device, whether the submission was sent to the appropriate FDA Center, and whether the device type is eligible for the 510(k) pathway.
submission must also meet the applicable processing requirements, including payment of the required user fee and receipt of a valid eCopy under the guidance. If the submission is incomplete, FDA may issue an RTA designation identifying the missing elements.
Applying the Acceptance Checklist
The acceptance checklist is a central part of the RTA process. FDA provides separate checklists for Traditional, Abbreviated, and Special 510(k) submissions. Manufacturers should use the checklist corresponding to their submission type to verify that required information has been addressed.
Checklist items may cover:
- Administrative and organizational elements.
- Device description, intended use and indications for use.
- Proposed labeling and instructions for use.
- Materials and technological characteristics.
- Performance testing and supporting reports.
- Biocompatibility, sterilization and software information, where applicable.
- Clinical information, where applicable.
- Device-specific guidance, special controls and applicable standards.
Not every category applies to every device. However, manufacturers should explicitly address applicability and provide a rationale for omissions or deviations where appropriate. The presence of a test report does not automatically mean that its results are scientifically adequate; that evaluation belongs to substantive review.
Handling an RTA Notification
An RTA designation means the submission has not been accepted for substantive review because one or more required acceptance elements are missing or inadequately addressed at the administrative-completeness level.
When FDA issues an RTA notification, the manufacturer should review the completed checklist, identify the missing elements, and submit the requested information under the originally assigned 510(k) number. A new submission and user fee are generally not required solely to provide the missing RTA information.
FDA then conducts another acceptance review, generally within 15 calendar days of receiving the additional information. If a complete response is not received within 180 days of the RTA notification, FDA considers the 510(k) withdrawn and closes the submission.
Acceptance Review Versus Substantive Review
The acceptance review checks whether a 510(k) submission is administratively complete using the applicable FDA checklist, with a target completion time of 15 calendar days. The substantive review evaluates the scientific and technical evidence to determine whether the device is substantially equivalent to a legally marketed predicate. Acceptance does not mean FDA has cleared the device; it only allows the submission to proceed to substantive review.
Q-Submission Interaction
Before submitting a 510(k), manufacturers may interact with FDA through the Q-Submission Program, particularly when the device raises novel regulatory or scientific questions or when the manufacturer is unfamiliar with the review process.
Early interaction can help clarify expectations for testing, regulatory strategy, and submission content. However, Q-Submission interaction does not replace the 510(k) acceptance checklist or guarantee acceptance or clearance.
Manufacturers should consult FDA’s separate guidance, Requests for Feedback and Meetings for Medical Device Submissions: The Q-Submission Program, for details on the available interaction mechanisms.
Maintaining Submission Readiness
RTA prevention should be treated as a structured regulatory quality activity rather than a final administrative check. Manufacturers should verify that all required sections are present, information is consistent throughout the submission, applicable checklist questions are addressed, and any omitted information is supported by a reasoned explanation.
Device-specific requirements should be considered carefully. For example, a device with patient-contacting components may require a biocompatibility assessment, while a sterile device may require information addressing sterilization. The relevant information depends on the device’s characteristics and applicable requirements.
Conclusion:
The FDA’s RTA policy provides a structured approach for checking the administrative completeness of 510(k) submissions before substantive review. It emphasizes appropriate submission selection, complete device and labeling information, applicable performance data, checklist based verification, and justified explanations for non applicable items or alternative approaches.
A submission that meets the acceptance criteria can proceed to substantive review, where FDA evaluates whether the device is substantially equivalent to a legally marketed predicate. If deficiencies lead to an RTA designation, the manufacturer must address the identified omissions and submit the missing information. Therefore, effective RTA management depends on early regulatory planning, systematic checklist review, and clear, well organized documentation.
References:
Refuse to Accept Policy for 510(k)s